
 Download Mendeley
Hydroxamate-based inhibitors reveal structural determinants of selectivity for Plasmodium falciparum aminopeptidase P.
Mills, B.J., Gregory, K.S., Cozier, G.E., Taweechai, S., Loughlan, E., McConkey, G.A., Isaac, R.E., Foster, R., Acharya, K.R.(2026) J Biol Chem 302: 111372-111372
- PubMed: 41850401 Search on PubMedSearch on PubMed Central
- DOI: https://doi.org/10.1016/j.jbc.2026.111372
- Primary Citation Related Structures:
9T1X, 9T1Y - PubMed Abstract:
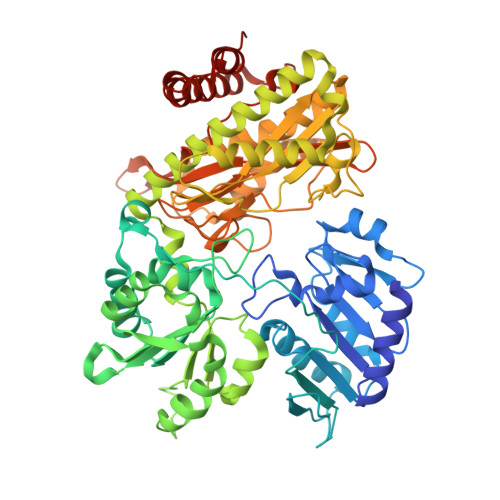
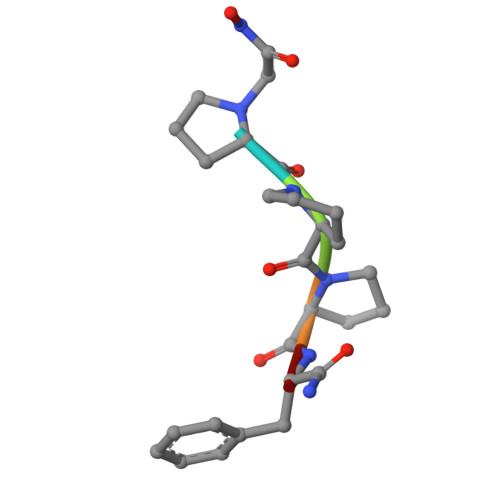
The malarial parasite, Plasmodium falciparum (Pf), utilizes aminopeptidases in the breakdown of hemoglobin-derived oligopeptides to release amino acids for protein synthesis during growth and asexual reproduction of erythrocytic stages of the parasite. However, a N-terminal peptide bond that involves proline is difficult to hydrolyze. Aminopeptidase P (APP) is capable of cleaving peptide bonds with proline in the second position. Inhibition of PfAPP is therefore an attractive strategy for developing therapeutics for the treatment of malaria by limiting the supply of amino acids at the erythrocytic stage. We employed the structure-activity relationship of an existing APP inhibitor, apstatin, to design a more potent PfAPP inhibitor by introducing a hydroxamic acid metal-binding group in place of the amino-alcohol of apstatin and an aromatic P4' moiety. A hydroxamic tetrapeptide with phenylalanine at P4' (6d) greatly increased the inhibitory potency (apstatin K i , 16 μM; 6d, K i 685 nM). Replacing the P3' proline of 6d with a 2-substituted piperidine (6e) further improved the potency (K i , 24 nM). Crystal structure analysis of PfAPP in complex with 6d and 6e showed binding at the active site with coordination of the hydroxamic acid metal binding group to the di-metal center, and several protein-inhibitor interactions involving domains II and III. A comparison of PfAPP-6e with human APP1 indicated that the P4' phenylalanine drives inhibitor potency and selectivity toward PfAPP, by forming an interaction with Tyr617 of the adjacent monomer within the dimer. The details presented here should be useful for the future design of potent and selective PfAPP inhibitors.
- School of Chemistry, University of Leeds, Leeds, UK.
Organizational Affiliation: